Acta Phys. -Chim. Sin. ›› 2026, Vol. 42 ›› Issue (4): 100224.doi: 10.1016/j.actphy.2025.100224
• ARTICLE • Previous Articles
Machine learning potentials for property predictions of two-dimensional group-Ⅲ nitrides
Jian Cao1, Chang Liu1, Danling Wang1, Haichao Li1, Lina Xu1,*( ), Hongping Xiao1, Shaoqi Zhan2, Xiao He3,*(), Guoyong Fang1,*()
- 1 College of Chemistry and Materials Engineering, Wenzhou University, Wenzhou 325035, Zhejiang Province, China
2 Department of Chemistry - ?ngstr?m Laboratory, Uppsala University, Uppsala 75120, Sweden
3 Shanghai Engineering Research Center of Molecular Therapeutics and New Drug Development, Shanghai Frontiers Science Center of Molecule Intelligent Syntheses, School of Chemistry and Molecular Engineering, East China Normal University, Shanghai 200062, China
-
Received:2025-07-18Revised:2025-11-20Accepted:2025-11-23Published:2026-01-29 -
Contact:Email: xulina@wzu.edu.cn (Lina Xu)xiaohe@phy.ecnu.edu.cn (Xiao He)fanggy@wzu.edu.cn (Guoyong Fang)

Cite this article
Jian Cao, Chang Liu, Danling Wang, Haichao Li, Lina Xu, Hongping Xiao, Shaoqi Zhan, Xiao He, Guoyong Fang. Machine learning potentials for property predictions of two-dimensional group-Ⅲ nitrides[J]. Acta Phys. -Chim. Sin. 2026, 42(4), 100224. doi: 10.1016/j.actphy.2025.100224
share this article
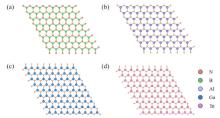
Fig 1
Optimized 2D structures of (a) h-BN, (b) h-AlN, (c) h-GaN and (d) h-InN."
Table 1
Experimental and calculated lattice parameter (Å) for h-BN, AlN, GaN and InN 2D monolayers."
| Compound | h-BN | h-AlN | h-GaN | h-InN | |||||||
| Parameters | Exp. | Cal. | Exp. | Cal. | Exp. | Cal. | Exp. | Cal. | |||
| Lattice parameter | 2.50–2.51 | 2.51 | 3.11–3.17 | 3.31 | 3.19 | 3.25 | 3.54 | 3.55 | |||
| Bond length | 1.44–1.45 | 1.45 | 1.80–1.83 | 1.80 | 1.82–1.85 | 1.87 | 2.15 | 2.09 | |||
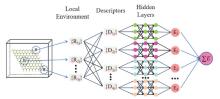
Fig 2
Structure of DP method."
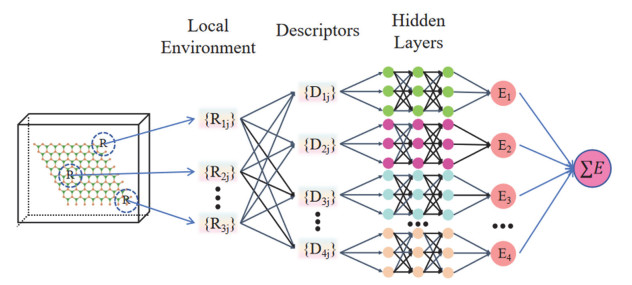
Fig 3
Schematic diagram of heat conduction direction."
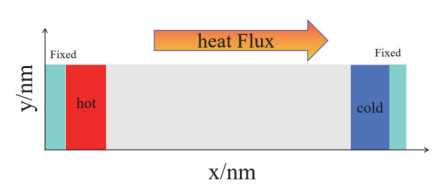
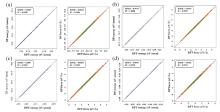
Fig 4
Comparison between DFT and DP results for four 2D Ⅲ-nitrides. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
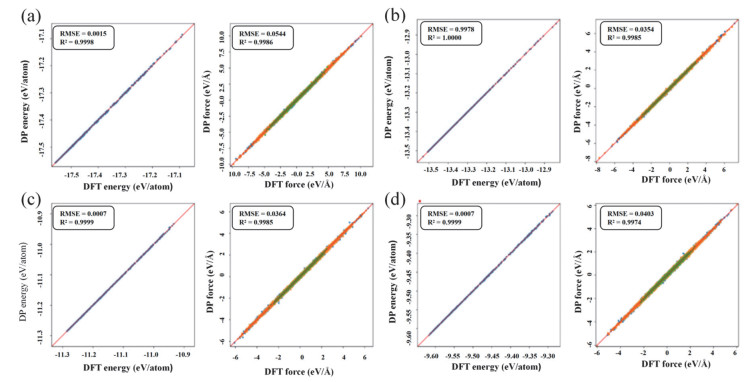
Fig 5
Phonon dispersion spectra of four 2D Ⅲ-nitrides via DFT and DP calculations. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
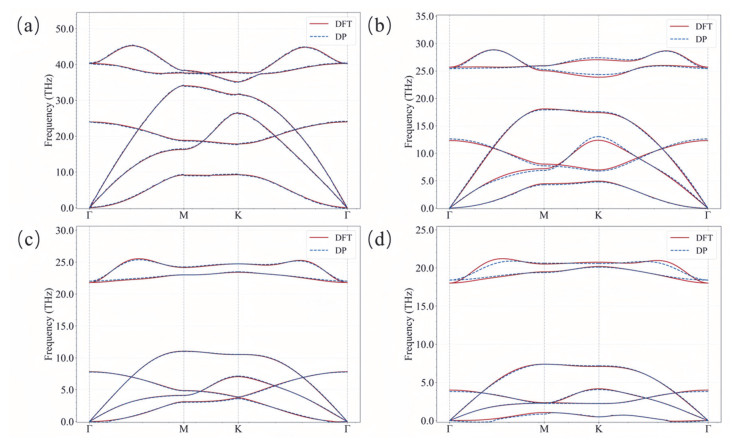
Fig 6
A of four 2D Ⅲ-nitride systems at different temperatures via DFT and DP calculations. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
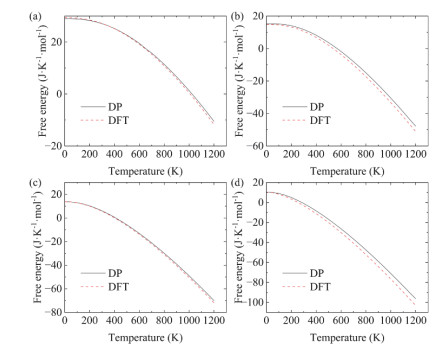
Fig 7
Cv of four 2D Ⅲ-nitrides at different temperatures via DFT and DP calculations. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
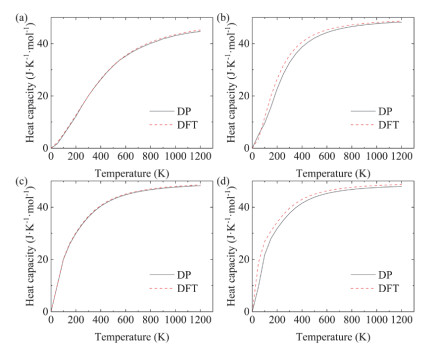
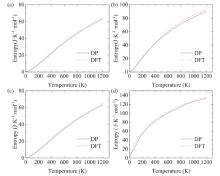
Fig 8
S of four 2D Ⅲ-nitrides at different temperatures via DFT and DP calculations. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
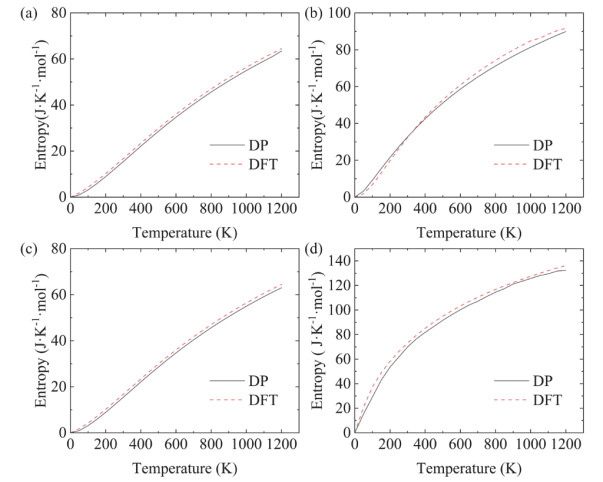
Fig 9
Uniaxial tensile stress-strain curves of 2D Ⅲ-nitrides along the x-direction. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
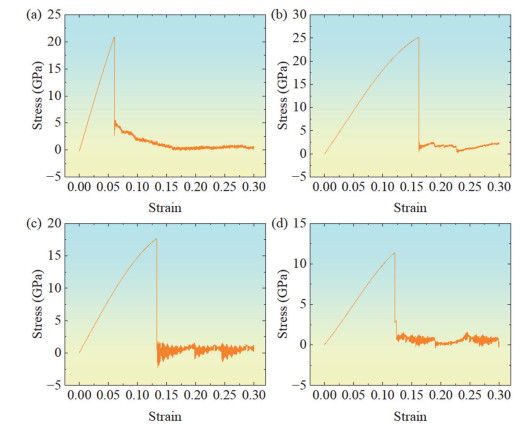
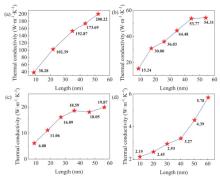
Fig 10
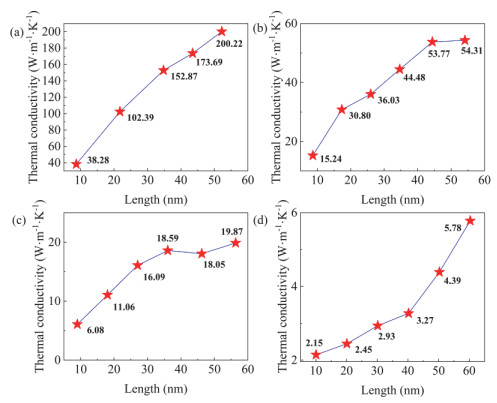
Length-dependent thermal conductivity of 2D Ⅲ-nitride materials extracted from NEMD simulations. (a) h-BN, (b) h-AlN, (c) h-GaN, and (d) h-InN."
| 1 |
F. Yang, L. Jin, L. Sun, X. Ren, X. Duan, H. Cheng. Adv. Mater. 2018, 30, 1801891.
doi: 10.1002/adma.201801891 |
| 2 |
S. Lv, S. Wang, L. Li, S. Xie, J. Yu, Y. Zhong, G. Wang, C. Liang, X. Xu, L. Zhang. Adv. Sci. 2023, 10, 2300780.
doi: 10.1002/advs.202300780 |
| 3 |
Y. Guo, H. Zhu, Q. Wang. ACS Appl. Mater. Interfaces 2018, 11, 1033.
doi: 10.1021/acsami.8b17341 |
| 4 |
S. Z. Butler, S. M. Hollen, L. Cao, Y. Cui, J. A. Gupta, H. R. Gutiérrez, T. F. Heinz, S. S. Hong, J. Huang, A. F. Ismach, et al.. ACS Nano 2013, 7, 2898.
doi: 10.1021/nn400280c |
| 5 |
A. H. Castro Neto, F. Guinea, N. M. R. Peres, K. S. Novoselov, A. K. Geim. Rev. Mod. Phys. 2009, 81, 109.
doi: 10.1002/adfm.202107499 |
| 6 |
D. Deng, K. S. Novoselov, Q. Fu, N. Zheng, Z. Tian, X. Bao. Nat. Nanotechnol. 2016, 11, 218.
doi: 10.1038/nnano.2015.340 |
| 7 |
A. Gupta, T. Sakthivel, S. Seal. Prog. Mater. Sci. 2015, 73, 44.
doi: 10.1016/j.pmatsci.2015.02.002 |
| 8 |
M. Xu, T. Liang, M. Shi, H. Chen. Chem. Rev. 2013, 113, 3766.
doi: 10.1021/cr300263a |
| 9 |
Q. Cui, G. Qin, W. Wang, K. R. Geethalakshmi, A. Du, Q. Sun. Appl. Surf. Sci. 2020, 500, 143993.
doi: 10.1016/j.apsusc.2019.143993 |
| 10 |
M. Q. Yang, N. Zhang, Y. J. Xu. ACS Appl. Mater. Interfaces 2013, 5, 1156.
doi: 10.1021/am3029798 |
| 11 |
K. Xia, V. I. Artyukhov, L. Sun, J. Zheng, L. Jiao, B. I. Yakobson, Y. Zhang. Nano Res. 2016, 9, 2182.
doi: 10.1007/s12274-016-1107-9 |
| 12 |
A. D. Oyedele, S. Yang, L. Liang, A. A. Puretzky, K. Wang, J. Zhang, P. R. Pudasaini, A. W. Ghosh, Z. Liu, C. M. Rouleau, et al.. J. Am. Chem. Soc. 2017, 139, 14090.
doi: 10.1021/jacs.7b04865 |
| 13 |
M. Corso, W. Auwa, M. Muntwiler, A. Tamai. Science 2004, 303, 217.
doi: 10.1126/science.1091979 |
| 14 |
J. Yu, L. Wang, Z. Hao, Y. Luo, C. Sun, J. Wang, Y. Han, B. Xiong, H. Li. Adv. Mater. 2020, 32, 1903407.
doi: 10.1002/adma.201903407 |
| 15 |
Z. Wang, G. Wang, X. Liu, S. Wang, T. Wang, S. Zhang, J. Yu, G. Zhao, L. Zhang. J. Mater. Chem. C 2021, 9, 17201.
doi: 10.1039/D1TC04022G |
| 16 |
B. Pécz, G. Nicotra, F. Giannazzo, R. Yakimova, A. Koos, A. Kakanakova-Georgieva. Adv. Mater. 2020, 22, 2006660.
doi: 10.1002/adma.202006660 |
| 17 |
I. A. Aleksandrov, T. V. Malin, K. S. Zhuravlev, S. V. Trubina, S. B. Erenburg, B. Pécz, Y. V. Lebiadok. Appl. Surf. Sci. 2020, 515, 146001.
doi: 10.1016/j.apsusc.2020.146001 |
| 18 |
M. E. Khan, M. Aamir, C. Ming, Y.-Y. Sun, Y.-H. Kim. FlatChem 2025, 51, 100846.
doi: 10.1016/j.flatc.2025.100846 |
| 19 |
E. Sandre, A. Pasturel. Mol. Simul. 1997, 20, 63.
doi: 10.1080/08927029708024168 |
| 20 |
R. Vink, G. Barkema, W. Van der Weg, N. Mousseau. J. Non-Cryst. Solids 2001, 282, 248.
doi: 10.1016/S0022-3093(01)00342-8 |
| 21 |
J. Tersoff. Phys. Rev. B 1988, 37, 6991.
doi: 10.1103/physrevb.37.6991 |
| 22 |
W. D. Cornell, P. Cieplak, C. I. Bayly, I. R. Gould, K. M. Merz Jr., D. M. Ferguson, D. C. Spellmeyer, T. Fox, J. W. Caldwell, P. A. Kollman. J. Am. Chem. Soc. 1995, 117, 5179.
doi: 10.1021/ja955032e |
| 23 |
L. D. Schuler, X. Daura, W. F. van Gunsteren. J. Comput. Chem. 2001, 22, 1205.
doi: 10.1002/jcc.1078 |
| 24 |
K. Wan, J. He, X. Shi. Adv. Mater. 2023, 36, 2305758.
doi: 10.1002/adma.202305758 |
| 25 |
D. Shvets, A. Gorbatov. Eur. Phys. J. Plus 2024, 139, 8685.
doi: 10.1140/epjp/s13360-024-05685-z |
| 26 |
J. Xia, Y. Zhang, B. Jiang. Chem. Soc. Rev. 2025, 54, 4790.
doi: 10.1039/d5cs00104h |
| 27 |
S. Hwang, J. Jung, C. Hong, W. Jeong, S. Kang, S. Han. J. Am. Chem. Soc. 2023, 145, 35.
doi: 10.1021/jacs.3c06210 |
| 28 |
J. Behler, M. Parrinello. Phys. Rev. Lett. 2007, 98, 146401.
doi: 10.1103/PhysRevLett.98.146401 |
| 29 |
A. P. Bartók, M. C. Payne, R. Kondor, G. Csányi. Phys. Rev. Lett. 2010, 104, 136403.
doi: 10.1103/PhysRevLett.104.136403 |
| 30 |
M. A. Caro. Phys. Rev. B 2019, 100, 024112.
doi: 10.1103/PhysRevB.100.024112 |
| 31 |
J. Byggmästar, K. Nordlund, F. Djurabekova. Phys. Rev. Mater. 2022, 6, 083801.
doi: 10.1103/PhysRevMaterials.6.083801 |
| 32 |
A. Thompson, L. Swiler, C. Trott, S. Foiles, G. Tucker. J. Comput. Phys. 2015, 285, 316.
doi: 10.1016/j.jcp.2014.12.018 |
| 33 |
I. S. Novikov, K. Gubaev, E. V. Podryabinkin, A. V. Shapeev. Mach. Learn. : Sci. Technol. 2020, 2, 025002.
doi: 10.1088/2632-2153/abc9fe |
| 34 |
L. Zhang, J. Han, H. Wang, R. Car, W. E. Phys. Rev. Lett. 2018, 120, 143001.
doi: 10.1103/physrevlett.120.143001 |
| 35 |
R. Drautz. Phys. Rev. B 2019, 23, 249901.
doi: 10.1103/PhysRevB.100.249901 |
| 36 |
H. Dong, X. Wu, Y. Shi, P. Ying, W. Zhou, K. Xu, T. Liang, S. Xiong, S. Chen, Y. Wang, et al.. J. Appl. Phys. 2024, 135, 161101.
doi: 10.1063/5.0200833 |
| 37 |
G. Kresse, J. Furthmüller. Phys. Rev. B 1996, 54, 11169.
doi: 10.1103/PhysRevB.54.11169 |
| 38 |
A. D. Becke. J. Chem. Phys. 1992, 96, 2155.
doi: 10.1063/1.462066 |
| 39 |
J. P. Perdew, K. Burke, M. Ernzerhof. Phys. Rev. Lett. 1996, 77, 3865.
doi: 10.1103/physrevlett.77.3865 |
| 40 |
J. Ben, X. Liu, C. Wang, Y. Zhang, Z. Shi, Y. Jia, S. Zhang, H. Zhang, W. Yu, D. Li, X. Sun. Adv. Mater. 2021, 32, 2006761.
doi: 10.1002/adma.202006761 |
| 41 |
M. A. Qaeed, K. Ibrahim, K. M. A. Saron, A. Salhin. Superlattices Microstruct. 2013, 64, 77.
doi: 10.1016/j.spmi.2013.08.015 |
| 42 |
A. Costales, M. A. Blanco, A. Martí. J. Am. Chem. Soc. 2002, 124, 4116.
doi: 10.1021/ja017380o |
| 43 |
K. He, X. Zhang, S. Ren, J. Sun, arXiv: 1512.03385, https://doi.org/10.48550/arXiv.1512.03385.
|
| 44 |
H. Wang, L. Zhang, J. Han, W. E. Comput. Phys. Commun. 2018, 228, 178.
doi: 10.1016/j.cpc.2018.03.016 |
| 45 |
Y. Du, C. Hao, Z. Meng, C. Wang, K. Peng, Y. Tian, W. Duan, L. Yang, P. Lin, S. Zhang. Comput. Mater. Sci. 2024, 242, 113078.
doi: 10.1016/j.commatsci.2024.113078 |
| 46 |
W. Yu, Z. Zhang, X. Wan, J. Su, Q. Gui, H. Guo, H.-x. Zhong, J. Robertson, Y. Guo. Chem. Mater. 2023, 35, 6651.
doi: 10.1021/acs.chemmater.3c00524 |
| 47 |
W. Liang, G. Lu, J. Yu. J. Mater. Sci. Technol. 2021, 75, 78.
doi: 10.1016/j.jmst.2020.09.040 |
| 48 |
D. J. Evans, B. L. Holian. J. Chem. Phys. 1985, 83, 406.
doi: 10.1063/1.449071 |
| 49 |
A. P. Thompson, H. M. Aktulga, R. Berger, D. S. Bolintineanu, W. M. Brown, P. S. Crozier, P. J. in't Veld, A. Kohlmeyer, S. G. Moore, T. D. Nguyen, et al.. Comput. Phys. Commun. 2022, 271, 108171.
doi: 10.1016/j.cpc.2021.108171 |
| 50 |
T. Gao, H. He, Y. Liu, Z. Bian, Q. Chen, Q. Xie, Y. Liang, Q. Xiao. Surf. Interfaces 2023, 39, 102983.
doi: 10.1016/j.surfin.2023.102983 |
| 51 |
K. Dey, S. Shahriar, M. A. R. Anan, P. Malakar, M. M. Rahman, M. M. Chowdhury. RSC Adv. 2024, 14, 24483.
doi: 10.1039/d4ra04814h |
| 52 |
B. Mortazavi, M. Makaremi, M. Shahrokhi, Z. Fan, T. Rabczuk. Carbon 2018, 137, 57.
doi: 10.1016/j.carbon.2018.04.090 |
| 53 |
F. Müller-Plathe. J. Chem. Phys. 1997, 106, 6082.
doi: 10.1063/1.473271 |
| 54 |
A. Chernatynskiy, S. R. Phillpot. Phys. Rev. B 2010, 82, 134301.
doi: 10.1103/physrevb.82.134301 |
| 55 |
M. Ozsipahi, S. Jean, A. Beskok, A. A. Wilson. Int. Commun. Heat Mass Transf. 2025, 163, 108658.
doi: 10.1016/j.icheatmasstransfer.2025.108658 |
| 56 |
Y. Yang, J. Ma, J. Yang, Y. Zhang. ACS Appl. Mater. Interfaces 2022, 14, 45742.
doi: 10.1021/acsami.2c14871 |
| 57 |
Z. Li, S. Xiong, C. Sievers, Y. Hu, Z. Fan, N. Wei, H. Bao, S. Chen, D. Donadio, T. Ala-Nissila. J. Chem. Phys. 2019, 151, 234105.
doi: 10.1063/1.5132543 |
| 58 |
L. Chaput, A. Togo, I. Tanaka, G. Hug. Phys. Rev. B 2011, 84, 094302.
doi: 10.1103/physrevb.84.094302 |
| 59 |
S. K. Achar, L. Zhang, J. K. Johnson. J. Phys. Chem. C 2021, 125, 14874.
doi: 10.1021/acs.jpcc.1c01411 |
| 60 |
J. Callaway. Phys. Rev. 1959, 113, 1046.
doi: 10.1103/PhysRev.113.1046 |
| 61 |
G. A. Slack. J. Phys. Chem. Solids 1973, 34, 321.
doi: 10.1016/0022-3697(73)90092-9 |
| 62 |
N. A. Sakharova, A. F. G. Pereira, J. M. Antunes. Nanomaterials 2024, 14, 1736.
doi: 10.3390/nano14211736 |
| [1] | Han-Zhi Luo, Qi-Ming Liang, Zi-Xing Guo, Xin-Tian Xie, Jin-Peng Tang, Tong Guan, Ye-Fei Li, Si-Cong Ma, Ying-Chen Xu, Zhen-Xiong Wang, Cheng Shang, Zhi-Pan Liu. LASPAI: AI-powered platform for the future atomic simulation [J]. Acta Phys. -Chim. Sin., 2026, 42(6): 100235-. |
| [2] | Jiahao Lu, Xin Ming, Yingjun Liu, Yuanyuan Hao, Peijuan Zhang, Songhan Shi, Yi Mao, Yue Yu, Shengying Cai, Zhen Xu, Chao Gao. High-Precision and Reliable Thermal Conductivity Measurement for Graphene Films Based on an Improved Steady-State Electric Heating Method [J]. Acta Phys. -Chim. Sin., 2025, 41(5): 100045-. |
| [3] | Xiaochen Zhang, Fei Yu, Jie Ma. Cutting-Edge Applications of Multi-Angle Numerical Simulations for Capacitive Deionization [J]. Acta Phys. -Chim. Sin., 2024, 40(11): 2311026-. |
| [4] | Cheng Ma, Xiangyu Dou, Zeyu Liu, Peilong Liao, Zhiyang Zhu, Kaerdun Liu, Jianbin Huang. Application and Mechanism of a Novel CO2-Oil Miscible Flooding Agent, CAA8-X [J]. Acta Phys. -Chim. Sin., 2022, 38(8): 2012019-. |
| [5] | Hongwei Yu, Shi Li, Jinlong Li, Shaohua Zhu, Chengzhen Sun. Interfacial Mass Transfer Characteristics and Molecular Mechanism of the Gas-Oil Miscibility Process in Gas Flooding [J]. Acta Phys. -Chim. Sin., 2022, 38(5): 2006061-. |
| [6] | Haoran Lu, Yaqing Wei, Run Long. Charge Localization Induced by Nanopore Defects in Monolayer Black Phosphorus for Suppressing Nonradiative Electron-Hole Recombination through Time-Domain Simulation [J]. Acta Phys. -Chim. Sin., 2022, 38(5): 2006064-. |
| [7] | Houfu Song, Feiyu Kang. Recent Progress on Thermal Conduction of Graphene [J]. Acta Phys. -Chim. Sin., 2022, 38(1): 2101013-. |
| [8] | Mengting Li, Xingqun Zheng, Li Li, Zidong Wei. Research Progress of Hydrogen Oxidation and Hydrogen Evolution Reaction Mechanism in Alkaline Media [J]. Acta Phys. -Chim. Sin., 2021, 37(9): 2007054-. |
| [9] | Wenqiong Chen, Yongji Guan, Jiao Zhang, Junjie Pei, Xiaoping Zhang, Youquan Deng. Atomistic Insight into Changes in the Vibrational Spectrum of Ionic Liquids under External Electric Field [J]. Acta Phys. -Chim. Sin., 2021, 37(10): 2001004-. |
| [10] | Chengzhen Sun, Runfeng Zhou, Bofeng Bai. Electrostatic Effect-based Selective Permeation Characteristics of Graphene Nanopores [J]. Acta Phys. -Chim. Sin., 2020, 36(11): 1911044-. |
| [11] | Zhaolong Chen, Peng Gao, Zhongfan Liu. Graphene-Based LED: from Principle to Devices [J]. Acta Physico-Chimica Sinica, 2020, 36(1): 1907004-. |
| [12] | Hai LIU,Yi LI,Zhaoxia MA,Zhixuan ZHOU,Junling LI,Yuanhang HE. Study on the Initial Decomposition Mechanism of Energetic Co-Crystal 2, 4, 6, 8, 10, 12-Hexanitro-2, 4, 6, 8, 10, 12-Hexaazaiso-Wurtzitane (CL-20)/1, 3, 5, 7-Tetranitro-1, 3, 5, 7-Tetrazacy-Clooctane (HMX) under a Steady Shock Wave [J]. Acta Physico-Chimica Sinica, 2019, 35(8): 858-867. |
| [13] | Tao ZHANG,Yunguang QIU,Qichao LUO,Xi CHENG,Lifen ZHAO,Xin YAN,Bo PENG,Hualiang JIANG,Huaiyu YANG. Concentration Dependent Effects of Ca2+ and Mg2+ on the Phosphatidylethanolamine-Phosphatidylglycerol Bilayer [J]. Acta Physico-Chimica Sinica, 2019, 35(8): 840-849. |
| [14] | Xiejun XU,Xingqing XIAO,Shouhong XU,Honglai LIU. Computational Study of Thermosensitivity of Liposomes Modulated by Leucine Zipper-Structured Lipopeptides [J]. Acta Physico-Chimica Sinica, 2019, 35(6): 598-606. |
| [15] | Kangjie LYU,Yanqiu PENG,Li XIAO,Juntao LU,Lin ZHUANG. Atomistic Understanding of the Peculiar Dissolution Behavior of Alkaline Polymer Electrolytes in Alcohol/Water Mixed Solvents [J]. Acta Phys. -Chim. Sin., 2019, 35(4): 378-384. |
|
||